Researchers used AI to create variants of a CRISPR-Cas12-like nuclease, some of which show increased editing activity
Structure and evolution-guided design of minimal RNA-guided nucleases
The design of RNA-guided nucleases with properties not limited by evolution can expand programmable genome-editing capabilities. However, generating diverse multidomain proteins with robust enzymatic properties remains challenging. Here, we use a protein design strategy that couples a structure-guided inverse-folding model with evolution-informed residue constraints to generate active, divergent variants of TnpB, a minimal CRISPR-Cas12–like nuclease, termed SynTnpBs. High-throughput screening of artificial intelligence–generated variants yielded editors that retained or exceeded wild-type activity in bacterial, plant, and human cells. Cryo–electron microscopy–based structure determination of the most divergent variant revealed stabilizing contacts in the RNA–DNA interfaces across conformations, demonstrating the design potential of this approach. Together, these results establish a strategy for creating non-natural RNA-guided nucleases and conformationally active nucleic acid binders, enlarging the designable protein space. , Editor’s summary Enzyme design and engineering are challenging in part because there are few ways to improve catalytic activity but many ways to impair it. Evolution-aware approaches can leverage information from our collection of known natural sequences to guide the generation of diverse engineered enzymes that are more likely to retain function. Skopintsev et al . demonstrated engineering of an RNA-guided nuclease using inverse protein-folding models. Screening of candidates nominated by their approach revealed improvement of genome-editing activity. An experimental structure revealed how conformational dynamics are changed in the engineered enzymes, stabilizing the key RNA-DNA interface. —Michael A. Funk
Generative AI comes to gene editing
Profluent releases AI-designed gene editor, OpenCRISPR-1

BE4max and AncBE4max Are Efficient in Germline Conversion of C:G to T:A Base Pairs in Zebrafish
The ease of use and robustness of genome editing by CRISPR/Cas9 has led to successful use of gene knockout zebrafish for disease modeling. However, it still remains a challenge to precisely edit the zebrafish genome to create single-nucleotide substitutions, which account for ~60% of human disease-causing mutations. Recently developed base editing nucleases provide an excellent alternate to CRISPR/Cas9-mediated homology dependent repair for generation of zebrafish with point mutations. A new set of cytosine base editors, termed BE4max and AncBE4max, demonstrated improved base editing efficiency in mammalian cells but have not been evaluated in zebrafish. Therefore, we undertook this study to evaluate their efficiency in converting C:G to T:A base pairs in zebrafish by somatic and germline analysis using highly active sgRNAs to twist and ntl genes. Our data demonstrated that these improved BE4max set of plasmids provide desired base substitutions at similar efficiency and without any indels compared to the previously reported BE3 and Target-AID plasmids in zebrafish. Our data also showed that AncBE4max produces fewer incorrect and bystander edits, suggesting that it can be further improved by codon optimization of its components for use in zebrafish.

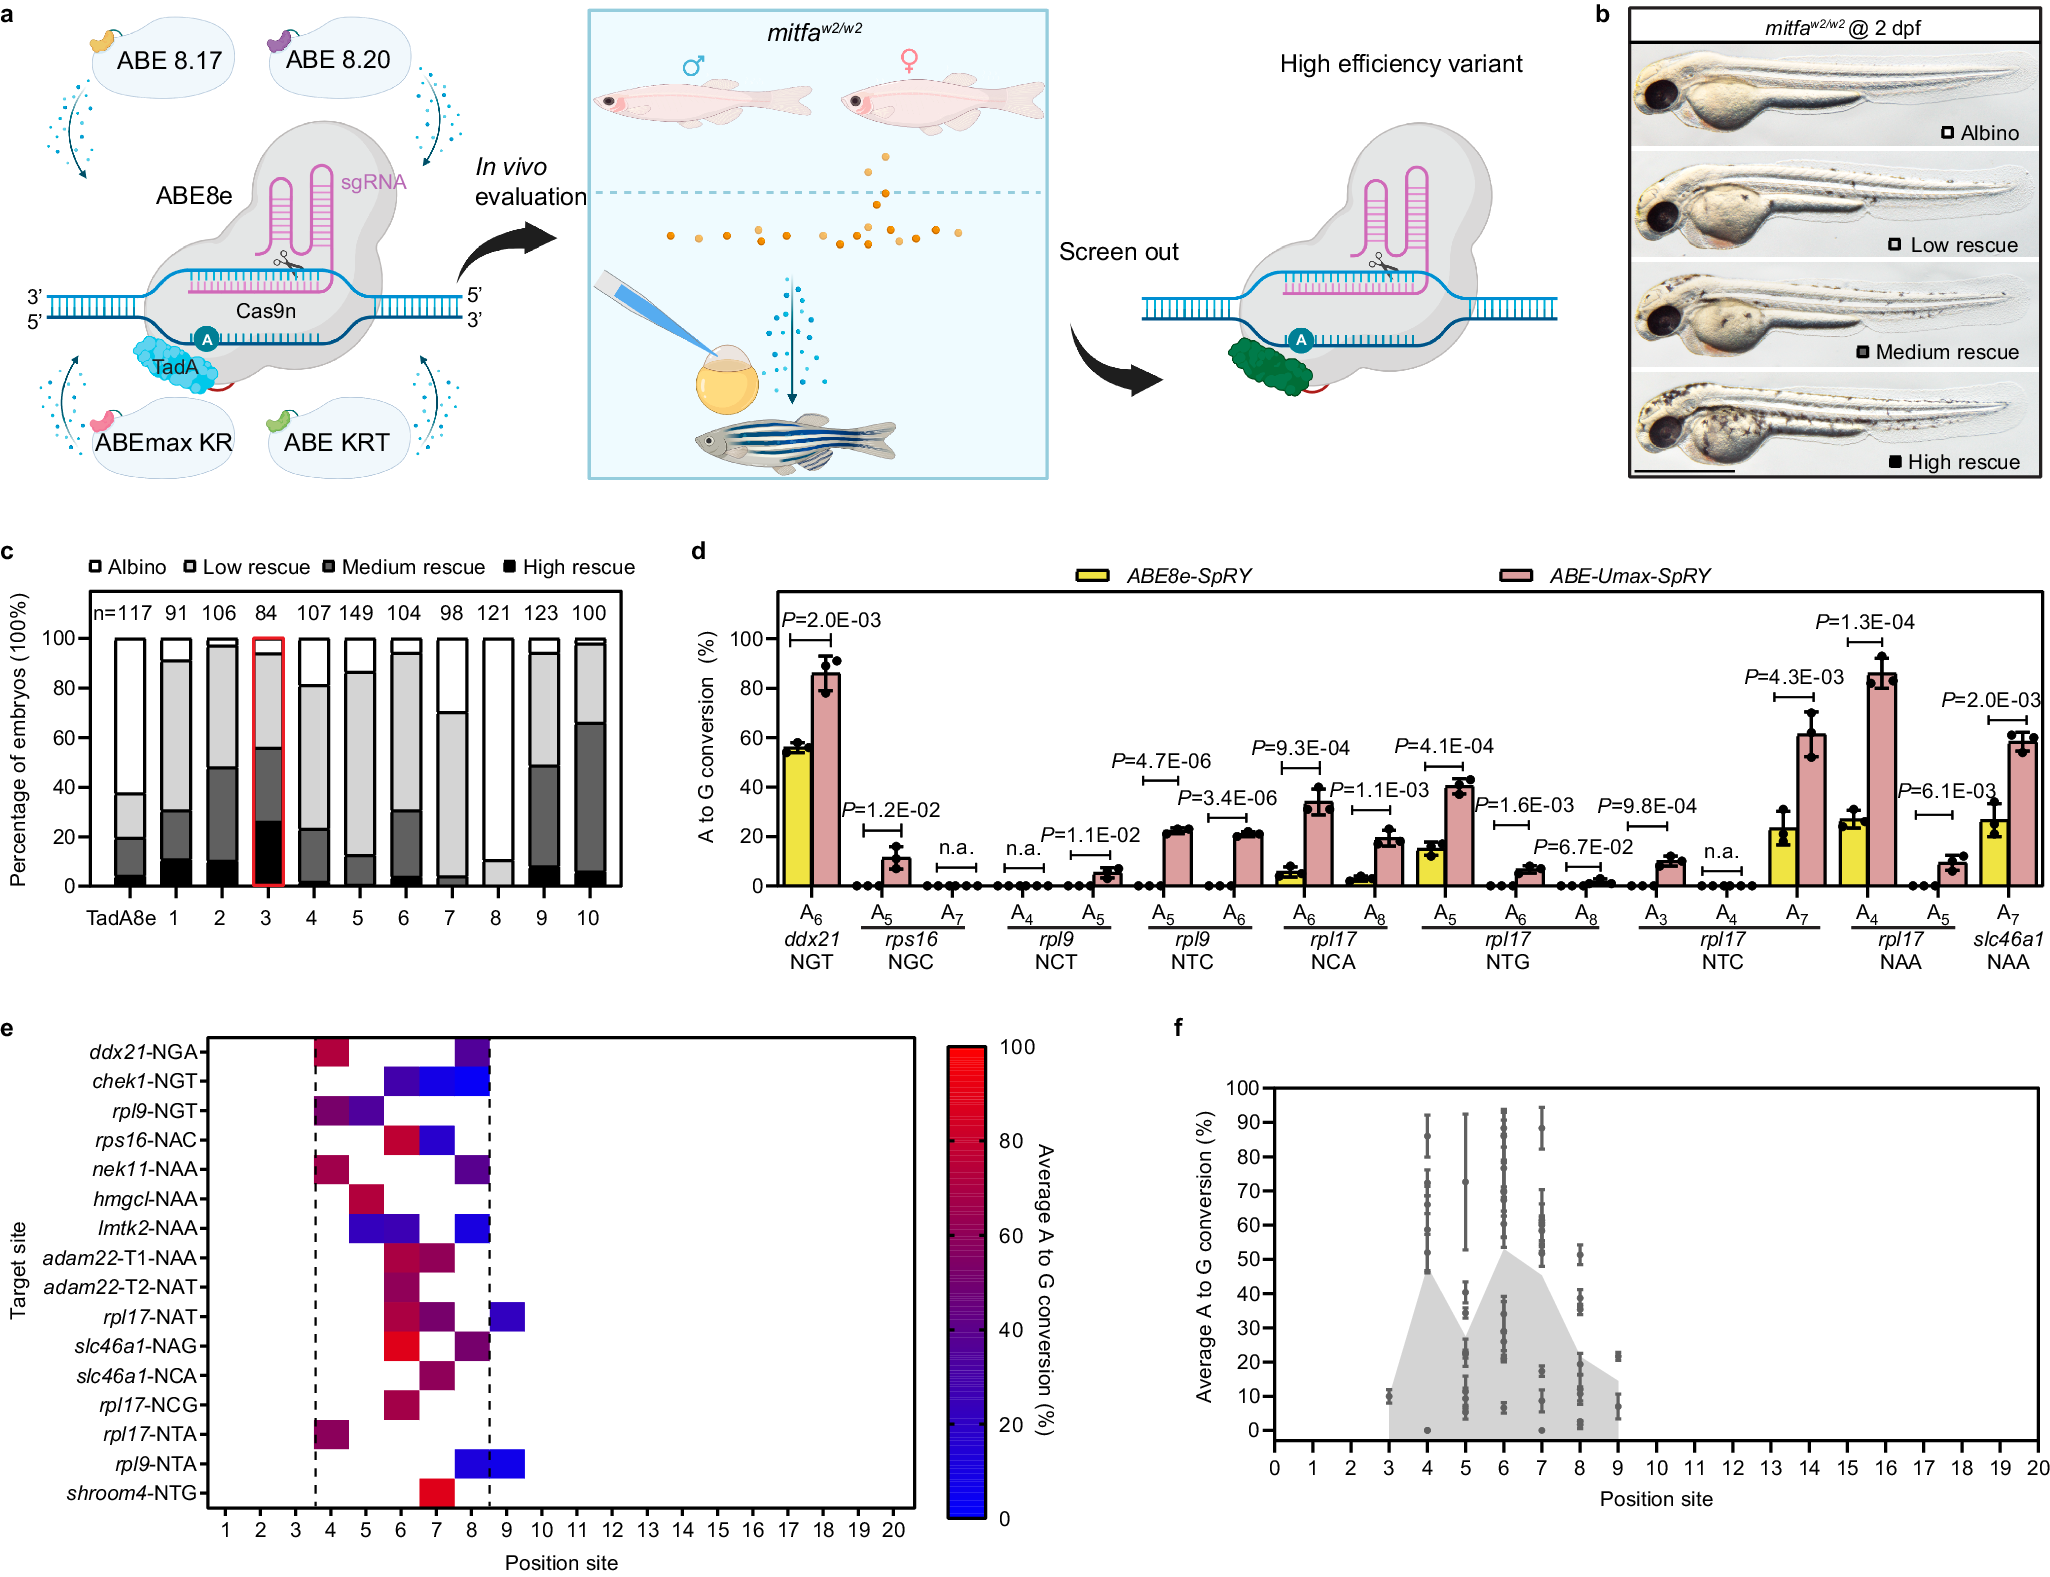
ABE-ultramax for high-efficiency biallelic adenine base editing in zebrafish
Advancements in CRISPR technology, particularly the development of base editors, revolutionize genetic variant research. When combined with model organisms like zebrafish, base editors significantly accelerate and refine in vivo analysis of genetic variations. However, base editors are restricted by protospacer adjacent motif (PAM) sequences and specific editing windows, hindering their applicability to a broad spectrum of genetic variants. Additionally, base editors can introduce unintended mutations and often exhibit reduced efficiency in living organisms compared to cultured cell lines. Here, we engineer a suite of adenine base editors (ABEs) called ABE-Ultramax (Umax), demonstrating high editing efficiency and low rates of insertions and deletions (indels) in zebrafish. The ABE-Umax suite of editors includes ABEs with shifted, narrowed, or broadened editing windows, reduced bystander mutation frequency, and highly flexible PAM sequence requirements. These advancements have the potential to address previous challenges in disease modeling and advance gene therapy applications.
How DNA Editing Could Change Life on Earth
Researchers hope to use the powerful biological tool to alter the genetic destiny of species and eliminate diseases.

Prime editing-installed suppressor tRNAs for disease-agnostic genome editing
Precise genome-editing technologies such as base editing1,2 and prime editing3 can correct most pathogenic gene variants, but their widespread clinical application is impeded by the need to develop new therapeutic agents for each mutation. For diseases that are caused by premature stop codons, suppressor tRNAs (sup-tRNAs) offer a more general strategy. Existing approaches to use sup-tRNAs therapeutically, however, require lifelong administration4,5 or show modest potency, necessitating potentially toxic overexpression. Here we present prime editing-mediated readthrough of premature termination codons (PERT), a strategy to rescue nonsense mutations in a disease-agnostic manner by using prime editing to permanently convert a dispensable endogenous tRNA into an optimized sup-tRNA. Iterative screening of thousands of variants of all 418 human tRNAs identified tRNAs with the strongest sup-tRNA potential. We optimized prime editing agents to install an engineered sup-tRNA at a single genomic locus without overexpression and observed efficient readthrough of premature termination codons and protein rescue in human cell models of Batten disease, Tay-Sachs disease and cystic fibrosis. In vivo delivery of a single prime editor that converts an endogenous mouse tRNA into a sup-tRNA extensively rescued disease pathology in a model of Hurler syndrome. PERT did not induce detected readthrough of natural stop codons or cause significant transcriptomic or proteomic changes. Our findings suggest the potential of disease-agnostic therapeutic genome-editing approaches that require only a single composition of matter to treat diverse genetic diseases.
(PDF) Highly efficient genome editing using oocyte-specific z cas9 transgenic zebrafish
PDF | Since its first application, CRISPR/Cas9 rapidly becomes a routine technique to perform genome editing in a variety of biological systems. To... | Find, read and cite all the research you need on ResearchGate

Dean W. Ball on Twitter / X
Ok two more:1. AI will enable the design of custom enzymes that can catalyze industrial processes and chemical manufacturing faster than existing methods. In some cases this will make existing chemicals cheap enough that it becomes possible to use them for totally new use… https://t.co/jL3bNm0s9w— Dean W. Ball (@deanwball) October 9, 2026
AI creates first synthetic viruses
Genomic language model has huge potential to redesign organisms such as bacteria

Highly efficient genome editing using oocyte-specific zcas9 transgenic zebrafish. Liu Y, Zhang C, Zhang Y, Lin S, Shi DL, Shao M
Highly efficient genome editing using oocyte-specific zcas9 transgenic zebrafish Since its first application to induce mutations in mammalian cells (Cong et al., 2013; Mali et al., 2013), CRISPR/Cas9 rapidly becomes a routine technique to perform genome editing in a variety of biological systems due to its facile, robust, and multiplexable features (Hwang et al., 2013; Guo et al., 2014; Wang et al., 2013). In every system, Cas9 and sgRNA should co-express to induce mutations. This is typically achieved by injection of the sgRNA mixed with Cas9 mRNA or protein into the fertilized eggs in zebrafish. As the in vivo translation of capped RNAs delay the presence of sufficient Cas9 protein, the mRNA injection produces more mosaic animals and shows relatively lower efficiency when compared to Cas9 protein injection (Burger et al., 2016). However, it is much more cost effective than using commercially available Cas9 protein, especially in the case of large-scale mutation screens. To further simplify this procedure, lower the cost and maintain an acceptable high genome editing efficiency in zebrafish, we set out to generate a transgenic line with stable and specific Cas9 expression in the egg. zpc (zp3b) promoter (zpc0.5) was reported to drive robust and specific GFP expression in the oocyte (Onichtchouk et al., 2003), we thus fused it with a zebrafish codon optimized cas9 (zcas9) followed by an SV40 3' UTR (Fig. 1A and File S1; Liu et al., 2014), and introduced this construct to zebrafish genome by Tol2 mediated transgenesis (Urasaki et al., 2006). To screen female F0 fish harboring this transgene, the F0 female founders were outcrossed with the wild-type male, the resulting eggs were injected with an sgRNA targeting slc45a2 (Moreno-Mateos et al., 2015), a gene essential for melanin synthesis, and assayed for their pigmentation phenotype. We found that the offspring of one founder showed a high proportion of pigmentation defect. Some of them exhibited a homozygous mutant like albino phenotype, suggesting a robust Cas9 expression in the egg and a very high efficient gene disruption. To examine the specificity of zcas9 expression in the transgenic line, the offspring (F1) of this F0 founder were raised to adulthood, and the ovaries of female F1 transgenic fish were dissected and subjected to in situ hybridization (ISH) employing the probe of zcas9. We found that the transcripts of zcas9 were specifically distributed in oocytes with a diameter of 40-200 µm, at around IB-II stages, while in WT embryos, no hybridization signals were detected (Fig. 1B, C). We did not detect zcas9 expression in stage IA oocytes or in larger ones after stage II. Accordingly, by RT-PCR, we also failed
Bending the Curve: How AI is Transforming Scientific Discovery
Discover how researchers use LLMs and specialized AI models like AlphaFold as complementary tools to accelerate innovation and expand the scientific frontier.

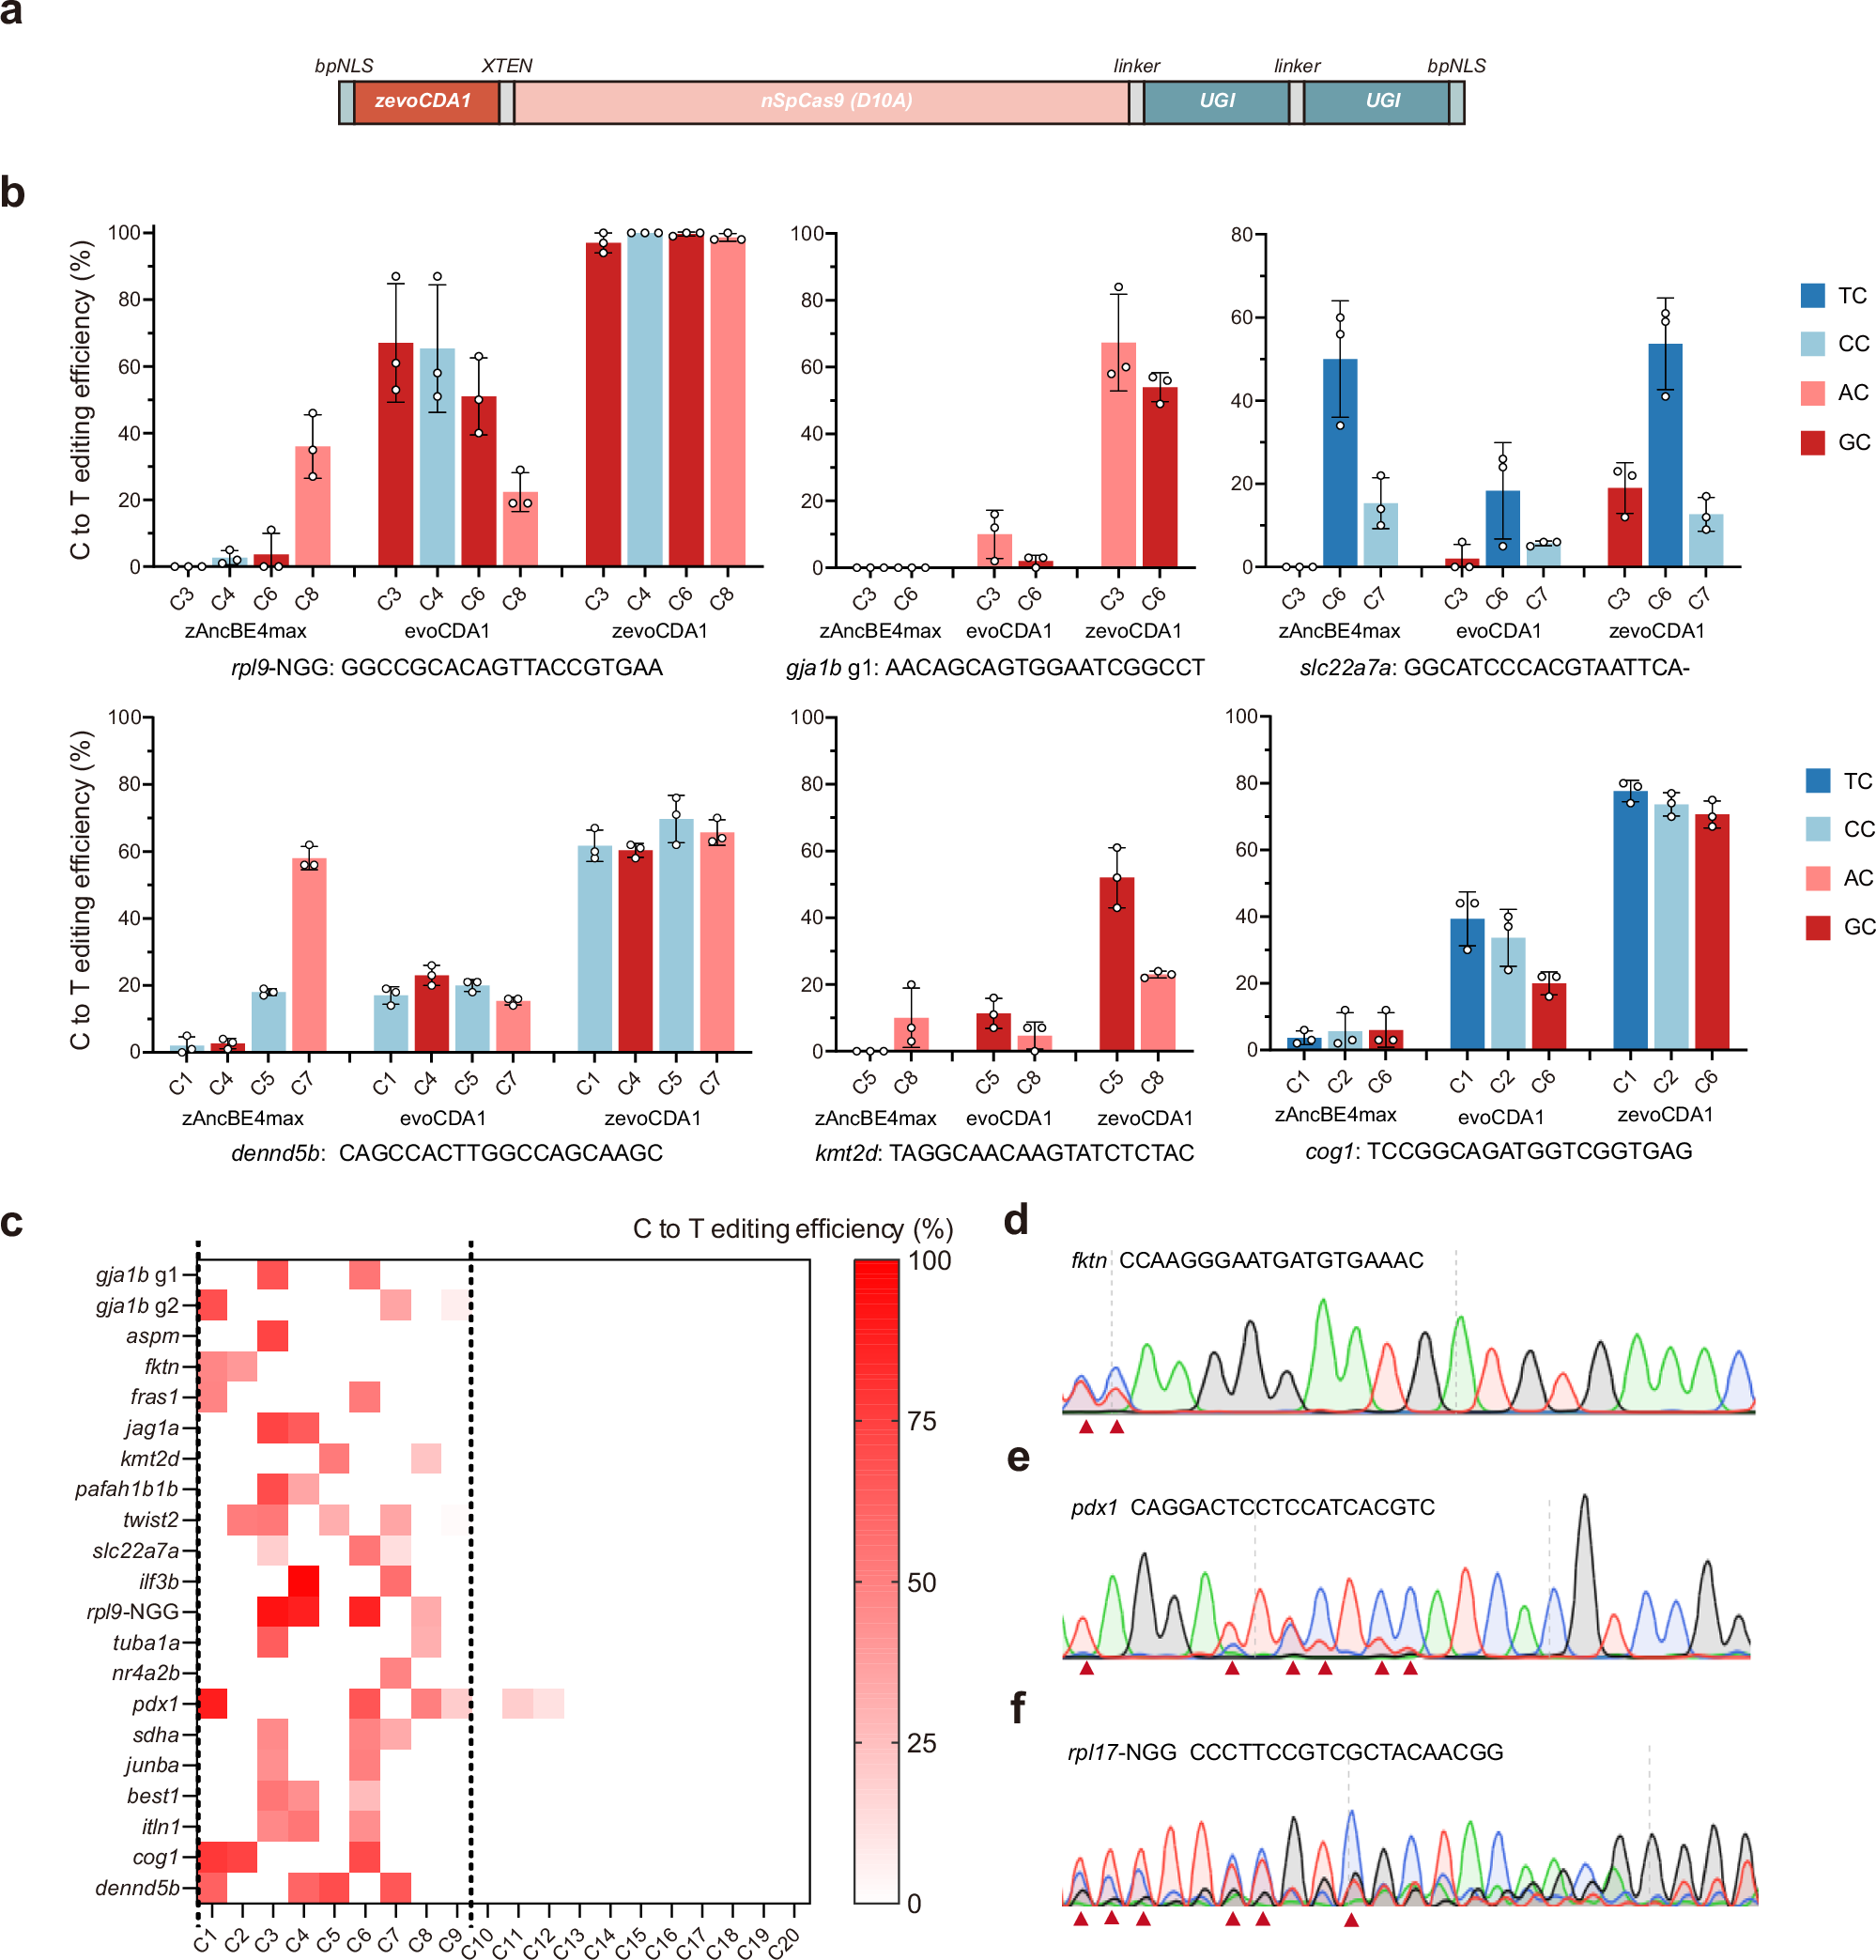
Cytosine base editors with increased PAM and deaminase motif flexibility for gene editing in zebrafish
Cytosine base editing is a powerful tool for making precise single nucleotide changes in cells and model organisms like zebrafish, which are valuable for studying human diseases. However, current base editors struggle to edit cytosines in certain DNA contexts, particularly those with GC and CC pairs, limiting their use in modelling disease-related mutations. Here we show the development of zevoCDA1, an optimized cytosine base editor for zebrafish that improves editing efficiency across various DNA contexts and reduces restrictions imposed by the protospacer adjacent motif. We also create zevoCDA1-198, a more precise editor with a narrower editing window of five nucleotides, minimizing off-target effects. Using these advanced tools, we successfully generate zebrafish models of diseases that were previously challenging to create due to sequence limitations. This work enhances the ability to introduce human pathogenic mutations in zebrafish, broadening the scope for genomic research with improved precision and efficiency.
No. 022: Palomar proof registry, autonomous protein design, synthetic consensus - The Hallway Track
The Hallway Track No. 022 - How AI is affecting the practice of science
Claude for Science. Thursday, November 12, at the Niels Bohr Institute, University of Copenhagen. In September, Anthropic reported that Claude agents found a CRISPR-like enzyme system in… | Jacob Langvad Nilsson | 17 comments
Claude for Science. Thursday, November 12, at the Niels Bohr Institute, University of Copenhagen. In September, Anthropic reported that Claude agents found a CRISPR-like enzyme system in bacteriophage DNA. Earlier in the same month, another AI lab said its model had settled a question about the Navier-Stokes equations, one of the seven Millennium Prize Problems. Results like these don't come from a lucky afternoon with a chatbot. They come from years of method development, careful engineering and building on the work that came before. What has changed is how much of that work a model can now take on, and the speed of which it does it. The latest models read the literature, write and run their own code, and work through a dataset for hours without losing track. In the enzyme case, hundreds of Claude agents searched DNA data for 21 hours and flagged a pattern nobody had asked them to look for. The method still has to be sound. But a researcher with a good question can now test far more ideas in a week than before. That is what we want to show. How researchers here set up their work so that stronger models can move it forward. The evening is about how the Danish science community uses Claude to drive new research and breakthroughs. Four STEM researchers from academia and industry each show one case. What they asked Claude, how they worked, what they found, and what failed. The room is part of it. Niels Bohr lectured in Auditorium A, and in the 1930s Heisenberg and Pauli sat in the front row. We will meet in this historical auditorium and get a glimpse of frontier scientific research using Claude. ▪️ Doors open 15:30, programme 16:00 to 19:00 ▪️ Four case studies from Danish research ▪️ 50 seats on the original wooden benches ▪️ Registration by approval Thanks to Daniel Murnane for making this happen. Thanks to the Niels Bohr Institute and the University of Copenhagen for having us. To Katrin Hjorth Thordardottir for logistics. And thanks to Anthropic and the Claude Community team, Martine N., Cassie Baquero Mayes, Chiara Massironi for your continued support. Link to sign up in the first comment below. We are still confirming speakers. Which researcher in Denmark should I talk to? | 17 comments on LinkedIn

Biohub releases a world model of protein biology
Open AI models used for a wide range of scientific applications, including accelerating the design of therapeutic molecules.